Download DS BIOVIA Materials Studio 2026

BIOVIA (Accelrys) Materials Studio 2026 is the latest release of BIOVIA’s comprehensive modeling and simulation environment for researchers in materials science and chemistry. Materials Studio enables researchers to predict and understand the relationships between the atomic and molecular structure of materials and their properties and behavior.
Materials Studio is an integrated modeling and simulation environment designed for constructing, manipulating, visualizing, simulating, and analyzing molecules, crystalline materials, surfaces, polymers, and mesoscale structures. It combines quantum, atomistic (classical), mesoscale, and statistical simulation methods, enabling researchers to investigate materials across different length and time scales. The platform also provides advanced tools for crystal structure analysis, crystal growth, and materials property prediction.
With its broad range of modeling and simulation capabilities, Materials Studio supports research and development in areas including catalysts, polymers, composites, metals, alloys, pharmaceuticals, batteries, semiconductors, photovoltaics, and other advanced materials.
Advantages of BIOVIA Materials Studio
Materials Studio provides an “in silico first” approach that allows researchers to investigate and optimize materials before conducting costly physical experiments.

DS BIOVIA Materials Studio 2024
BIOVIA Materials Studio
BIOVIA Materials Studio provides a comprehensive range of simulation and analysis tools covering quantum, atomistic, mesoscale, statistical, analytical, and crystallization modeling. By integrating multiple simulation approaches into a single platform, Materials Studio enables researchers to investigate materials across different length and time scales and gain deeper insights into material properties and behavior.
Kinetix
The Kinetix module uses the Kinetic Monte Carlo (KMC) method to provide a detailed understanding of the evolution of species concentrations and reaction rates on catalyst surfaces over time. It accounts for key surface processes, including diffusion, adsorption, surface reactions, and desorption.
The main capabilities include:
- Constant Conditions: Simulates changes in the concentrations of surface species and reaction rates under constant temperature and pressure conditions.
- Temperature Programmed: Simulates temperature-dependent surface processes using a defined initial temperature, final temperature, and heating rate.
- Programmed Potential: Models changes in surface species concentrations and reaction rates as a function of an applied electrochemical potential, such as in cyclic voltammetry experiments.
Cantera
The Cantera module is designed to solve chemical reaction-rate equations and predict the evolution of gas-phase and surface species in complex reaction systems. It can work with established reaction mechanisms and can also be combined with quantum-chemical calculations, such as those performed using DMol³.
Key capabilities include:
- Equilibrium: Calculates the composition of chemical species at thermodynamic equilibrium.
- CSTR: Simulates the time-dependent behavior of chemical species in a Continuous Stirred-Tank Reactor under specified flow rate, composition, temperature, and pressure conditions.
- 1D Flame: Performs one-dimensional flame and combustion simulations.
- Plug Flow Reactor: Simulates plug-flow reactor systems and predicts the composition of reactants and products, including catalyst surface coverage when surface reactions are included.
- TPD: Simulates temperature-programmed desorption processes to investigate the transfer of species between catalyst surfaces and the gas phase and to analyze changes in surface coverage and desorption rates with temperature.
- Reaction Mechanism Conversion: Provides a script for converting third-party reaction mechanism data into a format compatible with Cantera.
- Python Scripting: Supports Python scripting to extend and automate Cantera workflows.
Key Modules
CASTEP
CASTEP is a first-principles quantum-mechanical simulation package based on density functional theory (DFT). It can be used to investigate electronic structure, geometry optimization, phonons, mechanical properties, optical properties, spectroscopy, and other properties of crystalline materials.
DFTB+
DFTB+ provides a computationally efficient quantum-mechanical approach based on Density Functional Tight Binding. It is particularly useful for studying larger systems where conventional DFT calculations may be computationally expensive.
DMol³
DMol³ is a density functional theory (DFT) quantum-mechanical code designed for molecular and materials simulations. It can be used for geometry optimization, electronic structure, chemical reactions, adsorption, spectroscopy, and materials-property calculations.
Forcite
Forcite provides classical molecular mechanics and molecular dynamics capabilities for investigating the structure, dynamics, and properties of molecular and condensed-phase systems. It supports a wide range of force fields and can be used for geometry optimization, molecular dynamics, property calculations, and trajectory analysis.
GULP
GULP (General Utility Lattice Program) provides atomistic simulation capabilities for crystalline and molecular systems. It supports a wide range of force fields and can be used for geometry optimization, lattice dynamics, phonon calculations, defects, diffusion, thermal properties, and related materials simulations.
ONETEP
ONETEP is a linear-scaling density functional theory code designed to perform electronic-structure calculations on large-scale systems. It provides efficient simulations of systems containing thousands of atoms while maintaining first-principles accuracy.
Mesocite
Mesocite provides mesoscale simulation capabilities for investigating materials and soft-matter systems at length scales beyond conventional atomistic molecular dynamics. It can be applied to polymers, membranes, surfactants, composites, and other complex materials.
QMERA
QMERA combines quantum mechanical and molecular mechanical methods to study large systems in which a chemically important region requires quantum-mechanical treatment while the surrounding environment can be represented using molecular mechanics.
Visualizer
The Materials Studio Visualizer provides an integrated environment for building, editing, visualizing, analyzing, and preparing molecular and materials models. It supports visualization of structures, trajectories, simulation results, and calculated properties.
A Multiscale & Multiphysics Solution
BIOVIA Materials Studio provides an integrated multiscale modeling environment for investigating materials from the quantum and atomistic levels to mesoscale and statistical models.
Researchers can use Materials Studio to design, simulate, and analyze a wide variety of materials and chemical systems, including catalysts, pharmaceuticals, polymers, composites, alloys, batteries, semiconductors, and advanced electronic materials.
Materials Studio Applications
- Consumer Packaged Goods
- Materials Science
- Metal Alloy Design
- Battery Materials
- Catalyst Design
- Semiconductors & Sensors
- Visualization & Statistical Analysis
- Chemicals & Solvents
- Pharmaceutical Development
- Photovoltaics & Organic Electronics
- Polymer Composites.
New features in Materials Studio 2023
- Biovia materials studio 2023 for xps spectroscopy
- Biovia materials studio 2023 for chemical reactions
- Biovia materials studio 2023 for coarse grained simulations
- Biovia materials studio 2023 for metal alloy design
- The time-temperature-transformation (ttt) diagram protocol
- Quantum mechanics –more parameters! More properties (castep, dftb+ upgrade to version 22.1, onetep – upgrade to version 6.1.12)
- Enhancements to materialsscript
- Mesocite Dissipative Particle Dynamics (Dissipative Particle Dynamics (DPD) )
New Protocols in Materials Studio 2023
- A set of protocols that provide TURBOMOLE workflows
- through the Pipeline Pilot Protocols dialog are now available
- which provides DFT, xTB and Wavefunction methods
- Single-point energy calculation for all methods
- Geometry optimizations (using xTB, DFT or MP2)
- Transition state optimizations (using xTB or DFT)
- IR spectra / Vibrational frequencies (using DFT)
- Polarizabilities (using DFT)
- Optical spectra (using DFT)
Whats new in Materials Studio
- Quantum Mechanics – DFT Functionals:
- New CASTEP Functional!
- More DMol3 Functionals!
- Better Materials – Better Batteries- Metal Alloys – From Density Functional- Theory To Calphad Databases
- DFTB+ Parameterization:
- New Scripts!
- COSMO-RS COSMOBase:
- New Feature!
- Classical Simulations
- New Protocol!
- New Parameters!
- LARGE METALLIC SYSTEMS:
- New Feature!
- PERFORMANCE & USABILITY
- New Feature!
- New Protocol!
- PROPERTIES & ANALYSIS
- COOP/COHP analysis
- New Feature!
- Forcite & Mesocite
- New Feature!
- ONETEP
- QMERA
- QSAR
- NEW! Fitting valence forcefeld parameters using Pipeline
- NEW! Fitting non-bond forcefeld terms using using
- UPDATED! DFTB+: Creating parameters for DFTB+
- DMol3: Kinetics of a Diels-Alder reaction
- Forcite: Calculating the diffusivity of a gas in a polymer
- Pipeline Pilot Connector: Cross-linking polymers usin Pipeline Pilot
- Reflex: Structure solution of 4-nitrophenylhexylurethane using a close-contact penalty
- OTHER MATERIALS STUDIO 2020 HIGHLIGHTS
- The CASTEP user interface has been signifcantly modifed to reflect the current state of the server code and to aid usability
- GULP: A simulated annealing task has been added to GULP to provide flexible control over temperature in molecular dynamics simulations.
New in BIOVIA Materials Studio 2024
- Granular Dynamics – A new simulation method in Mesocite for modeling the motion of micron-scale spherical particles with negligible thermal velocity.
- Extended Mesocite Capabilities – New tasks including Anneal, Quench Dynamics, Confined Shear, Solvation Free Energy, Cohesive Energy Density, and Mechanical Properties.
- GPU-Accelerated DFTB+ – DFTB+ now supports the ELPA eigensolver with GPU acceleration and multiple-GPU support for improved performance on large systems.
- Effective Screening Medium in DMol³ – New ESM capabilities enable simulations of charged systems and electrochemical interfaces using vacuum or metal screening boundaries.
- DFT-D4 in CASTEP – CASTEP introduces the DFT-D4 dispersion correction for faster and accurate modeling of molecular crystals and van der Waals systems.
- Grand-Canonical ONETEP – ONETEP now supports grand-canonical simulations for systems with a variable number of electrons and a fixed external potential.
- New ReaxFF SEI2021 Library – GULP introduces a new ReaxFF library for modeling surface-electrolyte interphase (SEI) structures.
- New GULP Force Fields – Addition of the AA-CLP and Nitrates force-field libraries.
- ReaxFF with Electric Fields – GULP supports variable-charge calculations with charge derivatives and electric fields.
- Phonon Calculations with ReaxFF – ReaxFF calculations can include analytical second derivatives and phonons for evaluating phonon dispersion and phonon density of states.
- Enhanced MaterialsScript – All Forcite and Mesocite tasks available through the Visualizer can now be scripted using MaterialsScript.
- Bead Type Scripting – MaterialsScript now provides an interface for creating bead types for coarse-grained simulations.
- Improved Trajectory Analysis – Performance improvements for Rotational Time Correlation Function, VACF, Radial Distribution Function, and Mean Square Displacement analyses.
- Improved Forcite GPU Performance – Enhanced GPU performance, including support for calculations involving electric fields.
- New Uniaxial Barostat – Forcite dynamics introduces a new barostat option for simulations requiring uniaxial pressure control.
- Enhanced Reaction and Electrochemical Modeling – New capabilities support the modeling of complex chemical systems, electrochemical interfaces, and reactions under electric fields.
- Enhanced Visualizer – Improved pattern matching, close-contact analysis, force-field exclusion rules, and visualization workflows.
- Updated ONETEP – ONETEP is upgraded to academic version 6.1.45 with additional simulation capabilities.
- Updated GULP – GULP is upgraded to academic version 6.2 with new force fields, electric-field capabilities, and phonon functionality.
- Updated Intel MPI – Intel MPI is upgraded to version 2021.10 for improved parallel computing support.
- Updated Perl – Materials Studio 2024 includes Perl 5.36.0 across supported platforms.
New in BIOVIA Materials Studio 2025
- MACE Machine-Learned Forcefields – Introduction of MACE-OFF23 and MACE-MP-0 machine-learned forcefields in Forcite for high-accuracy molecular dynamics simulations.
- New Mesocite Builder – A dedicated builder for constructing mesoscale amorphous cells using bead-based mesomolecules and Mesocite forcefields.
- Advanced Mesoscale Model Building – New Construction, Packing, and ConfinedLayer tasks for streamlined mesoscale model generation.
- Mesoscale Conformation Randomization – New option to randomize molecular conformations when building mesoscale molecules.
- Bilayer Construction – New tools for building complex lipid and surfactant bilayer models, including single and double membranes.
- Martini 3 Support for Bilayer Modeling – Ready-made lipid, salt, and solvent molecules typed for the Martini 3 forcefield are available for membrane construction.
- Improved Amorphous Cell Performance – Faster construction of amorphous cells, particularly for large molecular systems.
- CASTEP D3 Dispersion Correction – Grimme D3 dispersion correction is available for selected GGA functionals, accelerating calculations involving nonlocal exchange-correlation functionals.
- CASTEP Piezoelectric Properties – Piezoelectric coefficients can now be calculated using the ultrasoft pseudopotential formalism.
- Improved Ultrasoft Pseudopotentials – New settings for automatically generated ultrasoft pseudopotentials for 4f and 5f elements.
- Elastic Anisotropy Analysis – New scalar log-Euclidian measures of elastic anisotropy are available from CASTEP and DMol³ elastic calculations.
- Ion Clustering Analysis – New Cluster Population Analysis tools for calculating and visualizing ion clustering in molecular dynamics trajectories.
- Heat Map Visualization – Generated heat maps provide an intuitive visualization of average ion-clustering behavior.
- Forcite GPU Enhancements – Improved GPU implementation, including Group Sum calculations for van der Waals and electrostatic interactions.
- Consistent NVIDIA GPU Performance – Improved Forcite GPU performance and consistency on NVIDIA graphics cards.
- Python 3.12.4 – Materials Studio 2025 includes an updated Python environment.
- Enhanced Mesocite Tutorials – New tutorials for constructing mesoscale amorphous cells and working with bead-based mesomolecules.
- MACE Forcefield Expansion in SP1 – Materials Studio 2025 SP1 adds MACE-MP-0b3 and MACE-MATPES-r2SCAN-0 for additional machine-learning-based simulation capabilities.
- Red Hat Enterprise Linux 9 Support – Materials Studio 2025 server gateways support newer Red Hat 9.x environments.
New in BIOVIA Materials Studio 2026
BIOVIA Materials Studio 2026 introduces new capabilities and enhancements across machine-learning force fields, quantum mechanical calculations, molecular dynamics, mesoscale modeling, scripting, and computational performance. The following features are among the major enhancements introduced in the 2026 release.
MACE Machine-Learned Force Fields
- MACE Fine Tuning: A new MACE Fine Tuning protocol allows users to refine MACE machine-learned force fields for specific materials and molecules.
- MACE Dispersion: Forcite now supports the D3 dispersion model with MACE force fields.
- Minimum Energy Path with FlexTS: The Forcite Minimum Energy Path task uses the FlexTS module to automatically calculate reaction pathways and energy barriers, with MACE machine-learned force fields supporting this functionality.
CASTEP
- Adaptive Compressed Exchange (ACE): CASTEP calculations using nonlocal or hybrid exchange-correlation functionals can benefit from significantly improved computational performance using ACE.
- Extended D3 and D4 Dispersion Corrections: CASTEP D3 and D4 dispersion correction schemes now include parameters for elements up to Z = 103.
- Infrared Spectra: IR spectra can now be calculated using ultrasoft pseudopotentials.
- Improved Convergence: Materials Studio can optionally attempt to improve calculation settings when CASTEP SCF or geometry optimization calculations fail to converge.
- Molecular Crystal Modeling: New workflows and tutorials support the modeling of molecular crystals using dispersion corrections.
DFTB+
- xTB Hamiltonians: DFTB+ now supports xTB Hamiltonians through the GFN1-xTB, GFN2-xTB, and IPEA1-xTB parameter sets.
- PTBP Parameter Set: A new Periodic Table Baseline Parameter (PTBP) set is available for solid-state modeling.
- GPU Acceleration: The ELPA eigenvalue solver is now the default on Linux servers and supports GPU acceleration.
ONETEP
- Improved Performance: ONETEP calculations can be optimized for speed rather than memory usage.
- Updated ONETEP Engine: The ONETEP solver has been updated to academic version 7.3.95, providing improved performance of the fast-density mode and reduced memory usage.
DMol³
- Improved Convergence: Materials Studio can optionally attempt to improve calculation settings when DMol³ SCF or geometry optimization calculations fail to converge.
- Enhanced Calculation Workflows: DMol³ continues to provide advanced density functional theory capabilities for electronic structure, geometry optimization, and materials-property calculations.
Forcite
- Minimum Energy Path: New Minimum Energy Path functionality provides automated reaction-pathway and energy-barrier calculations through FlexTS.
- MACE Support: Forcite supports MACE machine-learned force fields and D3 dispersion interactions.
- GPU Optimization: The GPU implementation of group-based sums has been optimized to improve computational performance.
- Viscosity Calculations: New scripts enable viscosity calculations using multiple molecular-dynamics trajectories and stress-autocorrelation analysis.
GULP
- Updated GULP Engine: GULP has been updated to academic version 6.4.
- Thermal Expansion: New capabilities include direct calculation of thermal expansion coefficients and anisotropic Grüneisen parameters.
- Enhanced ReaxFF: Additional ReaxFF capabilities include triple-bond and C2 special terms.
- Advanced Charge Equilibration: GULP now supports additional charge-equilibration schemes, including ACKS2, SCQEq, and EQEq, together with improved charge-derivative calculations.
- Improved QEq Control: Users have greater control over Coulomb interaction forms and the treatment of hydrogen atoms during charge equilibration.
COMPASS III
- Tetracoordinate Boron: COMPASS III now supports tetracoordinate boron compounds, including dialkylamino-substituted boranes.
Mesoscale Modeling
- Mesoscale Membrane Modeling: New workflows support the construction of mesoscale bilayer membranes in solution.
- Martini 3: Custom user-defined coarse-grained molecules based on Martini 3 can be incorporated into mesoscale membrane models.
- Improved Mesocite Workflows: Mesocite provides enhanced workflows for mesoscale modeling and membrane simulations.
CCDC ConQuest Integration
- CSD API Integration: Materials Studio CCDC ConQuest Search tasks now use the CSD API to search and extract structures from CCDC databases.
Python and MaterialsScript
- Python Scripting: Materials Studio now supports scripting with Python directly through MaterialsScript.
- Python and Perl: The MaterialsScript environment provides support for both Python and Perl scripting, allowing users to automate calculations and customize workflows.
- Updated Perl: Materials Studio 2026 includes Perl 5.40.2.
Other Improvements
-
- Improved Remote Job Management: Materials Studio can re-establish connections to remote jobs, including after restarting the client computer.
- Updated Installer: The Materials Studio installer is now provided as a .7z package.
- Improved GPU Support: Various GPU-related improvements have been implemented across the simulation modules.
- Enhanced Tutorials: New tutorials demonstrate workflows for molecular crystals, mesoscale membranes, and other advanced modeling applications.
Materials Studio 2019 details
- Version: 2019 Build 19.1.0.2353
- Operating system: 2019 Windows x64
- Crack: Full Crack
- Activation with: License (single user)
- Compressed file size: 1.95 GB
Materials Studio 2020.1 details
- Version: 2020 Build 20.1.0.5
- Operating system: windows x64 + Linux
- Crack: Full Crack
- Activation with: License (single user)
- Compressed file size: 1.83 GB + 720 MB
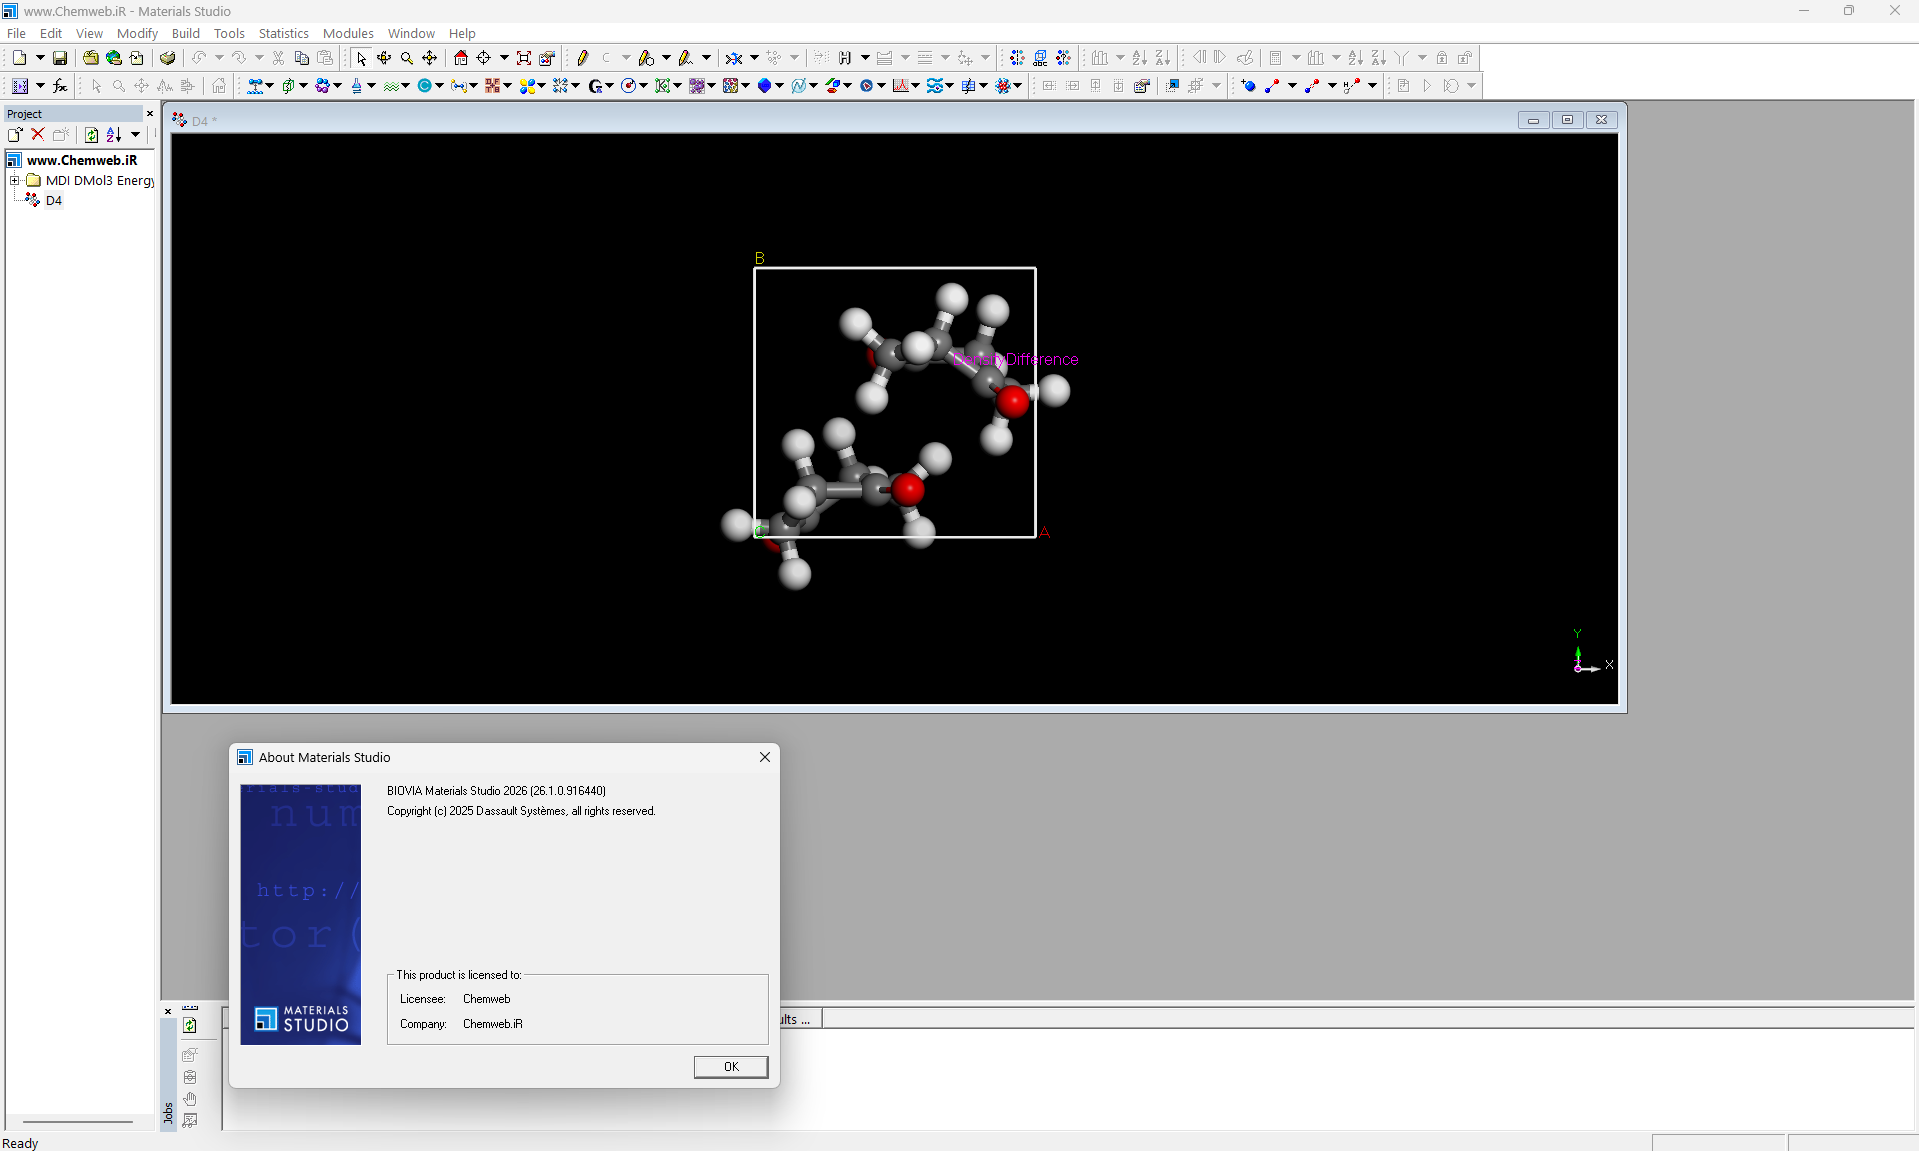
Download DS BIOVIA Materials Studio 2026
Materials Studio 2026 software details
- Version:2026 Build 26.1.0.916440 + 2023 Build 23.1.0.3829 + 2024 Build 24.1.0.321190
- Operating system: windows x64 + Linux
- Crack: Full Crack
- Activation with: License (single user)
- License validity: Permanent
- Compressed file size: 2.08 GB + 1.1 GB + 1.9 GB + 2.1 GB + 4.6 GB


